
A pharmaceutical equipment validation plan is not merely a collection of protocols prepared for an audit. It is the working blueprint that shows how a manufacturer will prove that equipment, utilities, software, and automated process controls consistently perform as intended. When the plan is thoughtful, it gives engineering, quality, production, and regulatory teams a shared view of what must be tested, why it matters, who owns each decision, and what evidence will be retained.
That evidence has a human purpose behind it. A minor inconsistency in mixing speed, dispensing weight, cleaning cycle, vacuum level, or label data may begin as a technical deviation, but it can ultimately affect dosage accuracy, product release, supply continuity, and patient safety. For pharmaceutical manufacturers, validation planning is therefore a practical exercise in controlling uncertainty before it reaches the market.
A well-built plan should cover the full equipment lifecycle rather than treating qualification as a one-time event. It should connect user needs, design decisions, risk controls, testing, data review, routine operation, maintenance, change management, and periodic assessment. The exact level of detail will vary between a laboratory balance, a clean-in-place skid, a tablet press, a sterile filling line, and a computerized batching system. The underlying principle, however, remains the same: demonstrate and maintain a state of control.
Start with intended use, not a generic validation checklist
The most reliable validation plans begin with a clear description of intended use. This sounds elementary, yet vague statements such as “the system will support production” create uncertainty later. The plan should explain what the equipment does, the materials it handles, the process step it supports, its operating environment, and the quality attributes it may influence.
For example, a vacuum system used in freeze-drying may affect chamber pressure, drying consistency, and product stability. An industrial weighing and batching system may control the quantity and sequence of active ingredients and excipients. An ultrasonic cleaning unit for pharmaceutical components may influence particulate and residue removal before assembly or filling. A laser marking or inkjet coding system may be central to lot traceability, packaging control, and anti-counterfeiting requirements.
Defining intended use also establishes the validation boundary. Teams should identify associated instruments, utilities, sensors, data interfaces, recipe management functions, cleaning equipment, alarms, and supporting software. Otherwise, a qualification effort may prove that the main machine runs while overlooking the process-support systems that determine whether it runs reliably and compliantly.
What a pharmaceutical equipment validation plan should include
Although document formats differ between organizations, a usable pharmaceutical equipment validation plan generally includes the following connected elements.
1. Scope, system description, and validation boundaries
The plan should name the equipment or system, identify its location and intended process, and distinguish what is included from what is excluded. It should map critical interfaces: utilities, compressed air, purified water, vacuum supply, environmental monitoring, manufacturing execution systems, laboratory information systems, and enterprise resource planning platforms where relevant.
This section should also state whether the work concerns a new installation, replacement equipment, a major upgrade, a transferred process, remediation, or periodic requalification. A replacement pump may seem like a maintenance task, for instance, but if its performance affects a critical vacuum process, the impact assessment may require more than a simple service record.
2. Roles, governance, and document control
Validation becomes fragile when responsibilities are assumed rather than assigned. The plan should identify accountable functions, commonly including engineering, manufacturing, quality assurance, quality control, automation, metrology, maintenance, validation, and external suppliers. It should define who prepares protocols, who executes tests, who reviews raw data, who approves deviations, and who gives final authorization for use.
Document control matters equally. The plan should reference governing procedures, approved templates, current equipment drawings, software versions, calibration records, standard operating procedures, and training requirements. Every protocol and report should be traceable to its approved version. In regulated operations, an impressive test result is of little value if the underlying document history is unclear.
3. Quality risk assessment
Risk assessment determines where validation effort belongs. Instead of testing every function with equal intensity, the team evaluates how equipment failure, incorrect operation, or data loss could affect patient safety, product quality, data integrity, and regulatory compliance.
A risk assessment may consider parameters such as temperature, pressure, mixing time, fill volume, weight tolerance, cleaning effectiveness, microbial control, barcode readability, audit-trail availability, and user access privileges. It should identify critical process parameters and critical quality attributes, then link them to specific controls and qualification tests.
The assessment should be living, not ceremonial. New supplier information, factory acceptance test results, installation discoveries, deviations, or process development findings may justify updates before final qualification. A plan that ignores emerging evidence is less useful than a shorter plan that responds to reality.
4. Requirements traceability
A requirements traceability matrix is one of the clearest ways to show that the validation program is complete. It links user requirements to functional specifications, design specifications, risk controls, test cases, acceptance criteria, deviations, and final outcomes.
Consider a computerized formulation system that must prevent an operator from dispensing an incorrect material. The user requirement may call for barcode-based material verification. The design specification may describe scanner integration and recipe logic. The test protocol should then challenge wrong-material scenarios, unreadable labels, interrupted connections, user override attempts, and audit-trail recording. Traceability makes it possible to see whether a requirement was simply documented or genuinely verified.
5. Qualification strategy and protocol sequence
The validation plan should describe the qualification approach, including which stages apply and how they relate to one another. A common structure includes design qualification (DQ), factory acceptance testing (FAT), site acceptance testing (SAT), installation qualification (IQ), operational qualification (OQ), and performance qualification (PQ). Not every system needs the same depth at each stage, but the rationale should be documented.
- DQ confirms that the proposed design meets intended use and applicable requirements.
- FAT and SAT can verify important functions before shipment and after installation, reducing late-stage surprises.
- IQ verifies correct installation, materials of construction, utilities, components, drawings, manuals, calibration status, and software configuration.
- OQ challenges operating ranges, alarms, interlocks, security functions, failure modes, and defined operating limits.
- PQ provides evidence that the integrated system performs consistently in routine or representative production conditions.
Protocol execution should be planned around realistic production circumstances. Testing only ideal conditions can leave important gaps. Where justified by risk, OQ should include boundary conditions, alarm verification, power interruption recovery, sensor failures, recipe changes, and operator access scenarios. PQ should reflect approved materials, trained operators, normal loading patterns, and controlled process conditions.
6. Predefined acceptance criteria and deviation handling
Acceptance criteria must be established before testing begins. They should be measurable, scientifically justified, and tied to the function being tested. “System operates satisfactorily” is not an acceptance criterion. More meaningful criteria define required ranges, accuracy, repeatability, response time, alarm behavior, reconciliation rules, or cleaning results.
The plan should also explain how deviations will be recorded, investigated, assessed for product or validation impact, corrected, and approved. A failed test does not automatically invalidate an entire project, but unexplained or casually closed deviations can weaken the evidence package. The important question is whether the deviation reveals a broader design, procedural, training, or control weakness.
How can process automation reduce formulation errors?
Process automation can reduce formulation errors by making critical steps less dependent on memory, manual transcription, visual judgment, and disconnected paper records. In pharmaceutical formulation, errors frequently arise at points where materials are selected, weighed, added, identified, mixed, transferred, or documented. Automation does not eliminate the need for qualified operators and sound procedures, but it can create dependable barriers against predictable mistakes.
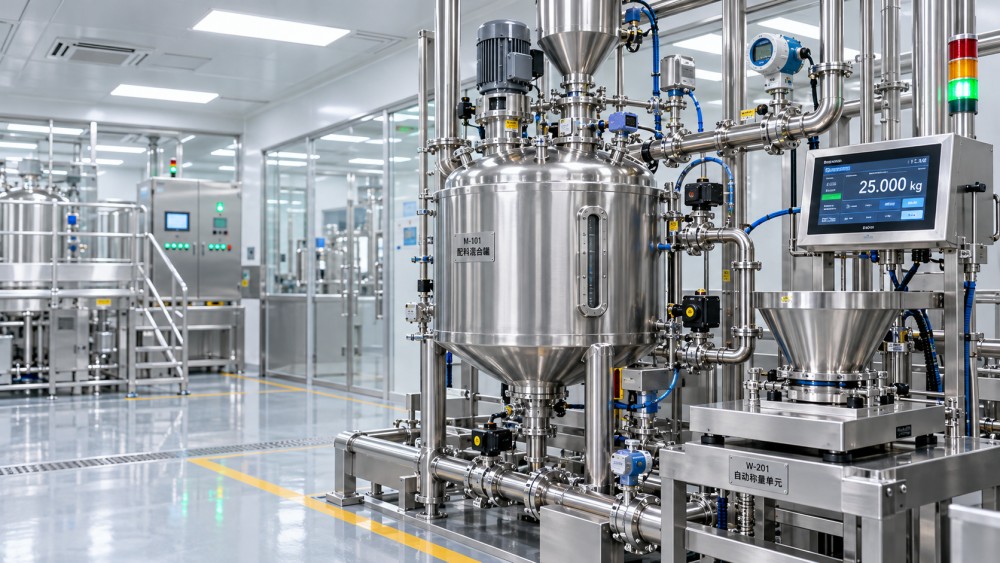
An automated weighing and batching system, for example, can guide the operator through the approved recipe sequence, verify raw-material identity through barcode or electronic confirmation, calculate target quantities, capture actual weights directly from calibrated scales, and prevent progression when tolerance limits are not met. This reduces the risk of selecting the wrong component, entering a number incorrectly, skipping a material, or using an outdated batch instruction.
For liquid and semi-solid formulations, automated controls can manage dosing rates, tank levels, agitation speed, mixing duration, temperature, and transfer status. Recipe-driven controls help ensure that the same approved parameters are applied from batch to batch. Integrated sensors can provide real-time feedback instead of relying solely on occasional manual checks. Where a parameter drifts, alarms and interlocks can prompt intervention before the deviation becomes a finished-product issue.
Automation also strengthens traceability. Electronic batch records, audit trails, time-stamped event logs, and controlled user access can reveal who performed an action, when it occurred, what value was recorded, and whether a recipe or setting was changed. These features are especially valuable when investigating a discrepancy or demonstrating compliance with data integrity expectations.
However, automation itself must be validated according to its impact. A highly automated process with poorly managed software access, uncontrolled configuration changes, inaccurate instruments, or unreviewed audit trails can introduce a different class of risk. The validation plan should therefore address computerized system functions, electronic records, user roles, backup and recovery, interfaces, alarm management, and cybersecurity controls where applicable.
Data integrity must be designed into the evidence trail
Validation records are only as credible as the data supporting them. The plan should specify how raw data will be generated, reviewed, stored, protected, and retrieved. This includes instrument printouts, electronic logs, calibration records, screenshots, sensor trends, test worksheets, calculation files, and any data transferred between systems.
Controls should support data that are attributable, legible, contemporaneous, original, and accurate, while also remaining complete, consistent, enduring, and available throughout the required retention period. In practical terms, that means unique user accounts rather than shared credentials, controlled time settings, reviewable audit trails, secure backups, defined access rights, and documented review procedures.
Teams should avoid treating data integrity as an IT-only topic. A scale that exports results into a spreadsheet, a coding system that receives lot information from a production database, or a vacuum controller that stores alarm histories can all create data-integrity considerations. The validation plan should identify these pathways early, before records begin to accumulate.
Plan for the period after approval
Equipment does not remain validated simply because a final report was signed. Wear, maintenance, software patches, new recipes, replacement parts, relocated sensors, revised cleaning agents, utility changes, and updated operating procedures can all affect the validated state. The plan should define how the organization will maintain control after release.
This lifecycle section should include calibration and preventive maintenance requirements, operator training, spare-parts controls where critical components are involved, periodic review, requalification triggers, and a formal change-control process. Change control should assess the proposed modification against intended use, risk assessment, validated ranges, documentation, and required testing. Small changes may require only documented justification; higher-impact changes may need partial or full requalification.
Periodic review is an opportunity to look beyond completed paperwork. Trend deviations, alarm frequency, maintenance history, out-of-tolerance events, rejected batches, audit-trail findings, and recurring operator interventions. These signals can show whether the system remains capable in actual use or whether its validation strategy needs attention.
Common weaknesses that make plans difficult to defend
Several recurring issues create avoidable pressure during inspections, internal audits, and deviation investigations. One is copying a previous validation plan without reassessing the new equipment’s intended use and risk profile. Another is allowing supplier documents to substitute for the manufacturer’s own acceptance criteria and quality oversight. Supplier testing can be valuable, but it should be reviewed in the context of the pharmaceutical process and approved requirements.
Other weak points include protocols written after testing has started, vague criteria, incomplete interface testing, missing calibration evidence, untested alarm conditions, and final reports that summarize success without addressing deviations. A particularly common gap is treating support equipment as peripheral. In reality, a dosing skid, vacuum pump, cleaning system, labeling device, or industrial balance may be the component that protects process consistency and traceability.
A stronger approach is to ask, at every stage: what could go wrong, how would we know, what prevents it, and where is the evidence? That question keeps the plan connected to the real factory floor rather than turning it into a compliance exercise detached from operations.
A validation plan should make routine confidence possible
The best pharmaceutical equipment validation plans are rigorous without becoming needlessly bureaucratic. They give teams a disciplined way to translate intended use into documented proof, focus testing where risk is highest, and preserve evidence that stands up to review. They also recognize that process-support technologies—precise weighing, controlled batching, validated cleaning, stable vacuum, reliable coding, and secure automation—often determine whether a manufacturing process stays within control.
When automation is thoughtfully specified and validated, it can reduce formulation errors while making each batch easier to understand, trace, and reproduce. That is the practical value of a validation plan: not simply proving that equipment worked on a test day, but building confidence that it will continue to support safe, consistent pharmaceutical production tomorrow.