
A batch record can look complete while the batch is already at risk. A scale may display a stable reading, the operator may follow the recipe, and the material may be released from the dispensing room—yet a small error in tare, unit selection, rounding, or material reconciliation can turn into a dosage-strength deviation later in the process. In pharmaceutical production, the concern is not simply whether a weight is “close enough.” It is whether the measured quantity remains within the approved process limits, can be traced to a calibrated instrument, and is supported by records that withstand review.
Pharmaceutical batching accuracy standards do not establish one universal tolerance for every material or batch. The acceptable range must be defined from the approved master batch record, product formulation, process validation knowledge, equipment capability, and applicable GMP requirements. A practical control system links three things: the allowable formulation tolerance, the measurement uncertainty of the weighing system, and documented evidence that the actual dispensing activity stayed under control.
Where batching deviations usually begin
The most visible deviation may be an incorrect active ingredient quantity, but not all risks are equally obvious. A low-dose active, potent compound, preservative, colorant, or processing aid can be highly sensitive to a small absolute error. By contrast, a large excipient charge may have a wider approved operating range but can still affect blend uniformity, granulation behavior, tablet hardness, dissolution, or fill weight when the error is significant relative to the formula.
Quality teams often see the problem first as a reconciliation discrepancy, an unexpected assay result, a failed in-process check, or an investigation triggered by an out-of-tolerance weighing event. By then, the original cause can be difficult to reconstruct. The deviation may have originated in one of several points:
- the balance was unsuitable for the target weight or minimum sample size;
- the approved tolerance was misunderstood as a generic percentage;
- the operator used the wrong unit, material status, container tare, or lot;
- the balance was calibrated but had not passed the required daily or shift verification;
- environmental effects such as airflow, vibration, static charge, or temperature instability affected the reading;
- the electronic batch record, manual worksheet, and actual dispensed quantity did not agree.
These failures are not solved by adding a final visual check. They require a weighing and dispensing process that prevents an unacceptable quantity from being accepted in the first place, identifies borderline conditions early, and preserves enough data for an independent reviewer to understand what happened.
Tolerance is a product and process decision, not a scale specification
A common misunderstanding is that a balance accuracy statement can be used as the product tolerance. It cannot. Balance readability, repeatability, linearity, eccentricity, calibration uncertainty, and minimum weight all describe aspects of the instrument’s measurement performance. The formulation tolerance defines how much variation in an ingredient quantity the approved process can accept. These are related, but they are not interchangeable.
For example, an approved batch record may specify a target quantity with a permitted range. That range should be scientifically justified through formulation development, process characterization, validation data, or other controlled product knowledge. It should reflect the ingredient’s function and the impact of variation on critical quality attributes. A tolerance that is appropriate for a bulk diluent may be unacceptable for a potent active or a material that strongly affects pH, viscosity, preservative effectiveness, or coating performance.
The measurement system must then be capable of controlling the approved range with sufficient margin. If the allowed range is narrow relative to the balance’s uncertainty or the practical variability of the dispensing operation, the process is vulnerable even when the scale is technically calibrated.
A defensible approach also distinguishes between process tolerance and action limits. The process tolerance is the approved acceptable range for the dispensed quantity. An internal action limit may be tighter, especially where a reading close to the edge of the approved range creates unnecessary risk. This gives operators and reviewers time to stop, evaluate, or reweigh before a true product specification issue occurs.
Choosing the right balance for the target weight
A balance that performs well at higher loads may not be suitable for a small dispense. Readability alone is not proof of suitability. A display that shows several decimal places can create a false impression of control when the repeatability, environmental conditions, and minimum usable weight do not support that precision.
The assessment should start with the smallest intended net quantity, not the largest vessel capacity. The smallest quantity is often the controlling case because measurement uncertainty becomes proportionally more significant at low weights. Where very small amounts are required, the site may need a more sensitive balance, a controlled pre-dilution approach, or a different approved dispensing strategy. The selected approach must preserve traceability and prevent calculation or transfer errors.
Before routine use, evaluate whether the balance performs acceptably across the expected working range. The assessment normally considers repeatability, linearity over relevant points, sensitivity, eccentric loading, and the balance location. A suitable test load should reflect real use as closely as possible. Testing only near the maximum capacity does not demonstrate capability for low-dose dispensing.
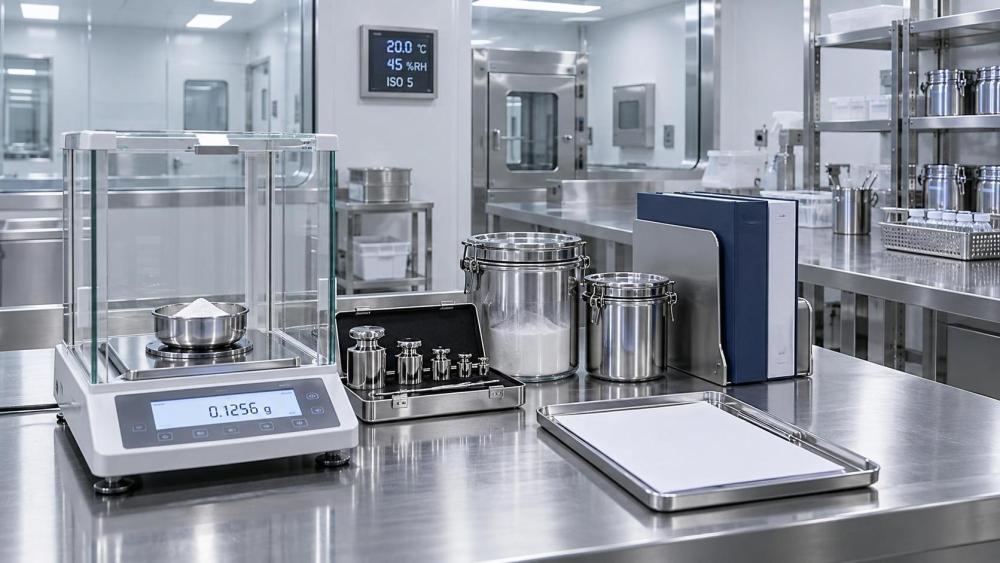
The balance environment matters as much as the balance model. Drafts from doors, HVAC outlets, or laminar airflow can destabilize readings. Static charge can cause powders, containers, and liners to behave unpredictably. Vibration from nearby equipment, uneven benches, poor leveling, and warm materials placed onto the pan can also affect results. A stable display is necessary, but operators should understand that stability indicators do not eliminate the need for controlled conditions.
Validation should prove the whole dispensing process
Calibration establishes traceability of the instrument against known standards at defined intervals. It does not, by itself, validate pharmaceutical batching. Process validation or qualification evidence must show that the overall dispensing and batching operation can consistently produce the intended result under normal operating conditions.
That broader evaluation may include the material identity check, container and liner tare process, transfer method, sequence of addition, operator instructions, barcode or electronic verification logic, reconciliation, and review process. A correctly weighed material can still be compromised if it is mislabeled, added to the wrong container, exposed to mix-up risk, or partially retained in a transfer container without proper accounting.
Define the intended operating range
Validation activities should not be built around an idealized center-point condition alone. The documented strategy should address the normal operating range: smallest and largest expected dispenses, different approved container types, representative material properties, expected shift patterns, and foreseeable environmental conditions. This does not mean creating artificial worst-case scenarios without justification. It means ensuring that the routine process, including its realistic sources of variation, has been considered.
Control software and data integrity
When batching is controlled through electronic systems, the software configuration becomes part of the accuracy control. The system should correctly identify materials, enforce units of measure, apply approved tolerances, retain the actual weight, record overrides, and maintain an audit trail. Interfaces between scales, manufacturing execution systems, warehouse systems, and batch-record platforms deserve particular attention. A correct balance reading can be converted, rounded, mapped, or transmitted incorrectly if interface controls are weak.
Any change affecting tolerances, formula quantities, balance type, software configuration, calibration method, material container, or dispensing workflow should be assessed through formal change control. The key question is whether the change could alter the ability to measure, verify, reconcile, or review the quantity accurately. Not every change requires full revalidation, but each should have a documented impact assessment and an appropriate level of testing.
Controls that make a batch record audit-ready
An auditor reviewing a batching event generally needs to follow a clear chain: approved instruction, approved material, qualified equipment, verified performance, actual result, review, and disposition of any exception. Gaps often arise because sites retain calibration certificates but cannot show the balance status at the time of use, or because they record an accepted final number without retaining evidence of a correction, override, or reweigh.
A practical batch record should make it possible to answer the following questions without relying on memory:
- Which material, lot, and status label were selected?
- Which balance or dispensing station was used, and was it within its approved calibration and verification status?
- What was the target quantity, approved tolerance, actual quantity, unit of measure, and container tare?
- Who performed the dispensing, and where required, who independently verified the activity?
- Were there any alarms, out-of-range readings, retries, overrides, spills, partial transfers, or adjustments?
- Did material reconciliation support the recorded quantity and final disposition?
Contemporaneous recording is essential. Writing the quantity later from memory, transcribing values from loose notes, or allowing unexplained corrections weakens data reliability. Paper records need controlled correction practices; electronic records need appropriate user access, audit trails, secure time-stamped entries, and review procedures. The aim is not administrative burden. It is to ensure that a critical manufacturing decision can be reconstructed accurately.
Handling a result near or outside the allowed range
A reading near the tolerance boundary should not automatically be treated as routine. First confirm that the displayed value is stable and that the correct unit, tare, material, and container are in use. Check the balance verification status and look for obvious environmental interference. Do not erase, overwrite, or casually repeat a questionable result simply to obtain a more convenient number.
When a value is outside the approved range, follow the approved deviation or exception procedure. The immediate priorities are to prevent unintended use of the material, preserve the original record, and determine whether the issue is an execution error, equipment concern, documentation error, or a legitimate process event requiring quality assessment. Reweighing may be appropriate only when the procedure permits it and the original result remains documented. Repeated measurements without a defined rationale can obscure the actual event and create data-integrity concerns.
The investigation should be proportionate to risk. A small discrepancy in a noncritical bulk material may require a different assessment from an error involving a potent active ingredient or a component with a narrow functional range. Review the material balance, scale checks, operator actions, audit trail, labels, and transfer steps before concluding that the issue was isolated. Where an equipment issue is suspected, evaluate potentially affected batches since the last known acceptable verification or calibration point according to the site’s documented quality system.
Frequent weak points in otherwise mature operations
One recurring weakness is applying a percentage tolerance to every component without considering low target weights. A percentage may produce an impractically narrow range for a small dispense or an overly broad range for a large one. Another is allowing manual rounding rules that differ between the batch record, balance display, and electronic system. Rounding must be defined so that it does not move an actual value across an acceptance boundary.
Sites also underestimate the importance of tare control. The container, liner, scoop, and residual powder can materially affect a low-weight dispense. Tare practices should be standardized, especially where disposable liners, nested containers, or moisture-sensitive materials are involved. A change in container type may require assessment even when the formula itself has not changed.
Finally, calibration labels should never substitute for operational discipline. Pharmaceutical batching accuracy standards are met when approved tolerances are scientifically justified, instruments are demonstrably fit for use, the work environment supports reliable measurement, and each dispensing event is recorded in a reviewable form. That combination protects dosage consistency while giving quality and safety teams a defensible basis for release decisions and audit response.