
On June 30, 2026, the U.S. FDA announced a new import screening pathway under its Advanced Manufacturing Import Fast Track (AMIFT) pilot, with Cleanroom Ultrasonic cleaning systems included in the first batch. From August 1, 2026, products in this category entering the U.S. market must be submitted with a cleanroom environmental test report issued by an ISO/IEC 17025-accredited laboratory under ISO 14644-1:2026. For manufacturers, exporters, import compliance teams, testing providers, and downstream buyers, the change is worth close attention because it connects customs clearance speed directly to documented cleanroom performance.
What the FDA Has Confirmed So Far
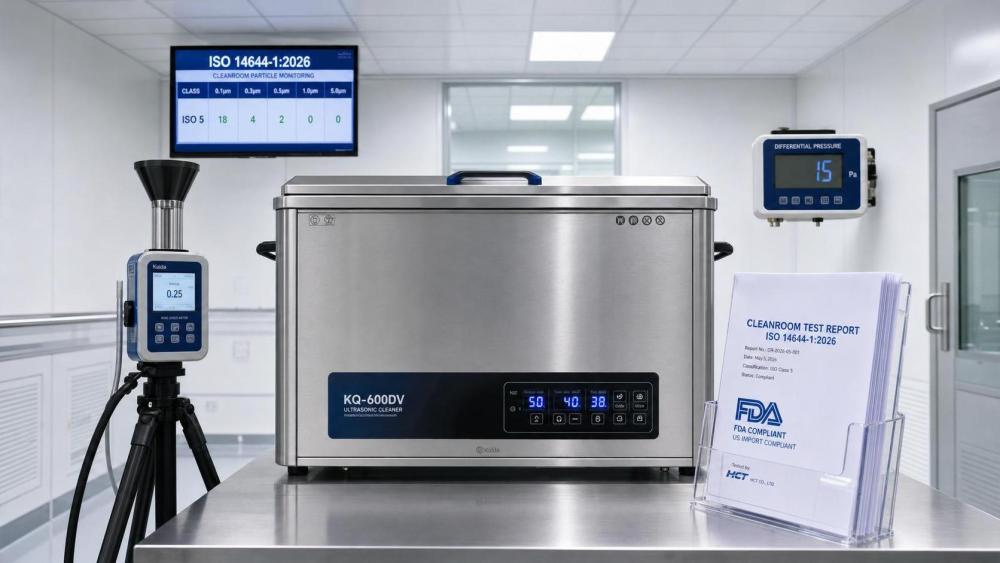
According to the provided event summary, the FDA launched the AMIFT pilot on June 30, 2026, and Cleanroom Ultrasonic cleaning systems are among the first covered products. Beginning August 1, 2026, all such equipment declared for entry into the U.S. must be accompanied by a cleanroom environmental measurement report based on ISO 14644-1:2026 and issued by an ISO/IEC 17025-accredited laboratory.
The report must include measured results covering Class 5/ISO 5 dynamic particle counts, airflow uniformity, and pressure differential gradient. The provided information also states that submissions failing to meet the requirement will trigger 100% on-site inspection and delayed release.
Where the Pressure Will Be Felt First
Export-facing equipment suppliers will face a documentation gate
From an industry perspective, manufacturers and direct trading companies shipping Cleanroom Ultrasonic systems to the U.S. may feel the immediate impact at the shipment preparation stage. The key issue is no longer only product declaration, but whether the required cleanroom report is complete, current, and aligned with the stated standard and accreditation requirement.
Compliance and import operations will need tighter file control
For importers, customs teams, and supply chain service providers, the practical impact is likely to appear in document review, filing timelines, and release planning. What deserves closer attention is that the stated consequence of non-compliance is not a minor procedural correction, but 100% on-site inspection and delayed release, which can affect delivery scheduling and internal coordination.
Testing and verification providers become part of the delivery timeline
Observably, laboratories and related service providers may become more central to transaction timing for this product category because the FDA requirement specifically points to ISO/IEC 17025-accredited laboratories and measured cleanroom data under ISO 14644-1:2026. For companies relying on external testing, laboratory capacity, report format, and turnaround time may become operational variables rather than background support work.
Buyers and end users may pay closer attention to readiness before shipment
Procurement teams and end users sourcing covered systems for the U.S. market may also be affected indirectly. Analysis shows that import readiness now depends in part on whether the supplier can provide the required report package before shipment, which may influence purchase confirmation, acceptance timing, and communication around delivery expectations.
What Companies Should Watch Right Now
Check whether current reports match the stated standard path
Companies involved in U.S.-bound shipments should first review whether existing cleanroom testing documents actually meet the requirement described in the event summary: ISO/IEC 17025-accredited laboratory issuance and ISO 14644-1:2026-based measurement content. A report that is technically related but not aligned with the stated framework may not support smooth filing.
Pay attention to the August 1, 2026 implementation date
The timeline matters because the rule is tied to market entry from a specific date. Businesses with orders close to that threshold may need to look carefully at filing schedules, shipment timing, and whether any in-process deliveries could fall into the new documentation window.
Separate policy wording from execution details
Analysis shows that the confirmed facts establish the reporting requirement and the consequence of non-compliance, but they do not fully describe every operational detail of enforcement. Companies should therefore distinguish between what is already explicit and what may still depend on later official clarification, especially in filing practice, review expectations, and documentation handling.
Prepare customer and supplier communication early
For businesses operating across manufacturing, export, logistics, and procurement, it would be practical to align internal and external communication now. The immediate focus should be on supplier qualification, report availability, document completeness, and contingency planning for possible inspection-related delays where compliance is uncertain.
Why This Looks Bigger Than a Single Filing Requirement
As an editorial observation, this update appears to be more than a narrow paperwork adjustment for one equipment segment. The FDA is linking a fast-track import framework with measurable cleanroom evidence for Cleanroom Ultrasonic systems, which signals closer scrutiny of documented manufacturing environment conditions at the point of market entry.
At the same time, it is more appropriate to understand this as an early-stage regulatory signal rather than a final picture of long-term market impact. The confirmed facts show a pilot structure, a defined product scope in the first batch, and a clear compliance trigger. What remains worth watching is whether similar documentation logic extends to additional advanced manufacturing equipment categories over time.
How the Market Should Read This Update
The immediate meaning of this development is straightforward: for covered Cleanroom Ultrasonic systems entering the U.S., cleanroom performance documentation is becoming an import clearance condition rather than an internal quality file alone. For the industry, the practical significance lies in the overlap between technical validation, trade compliance, and delivery certainty.
Taking a neutral view, this is best understood at present as a concrete short-term compliance change and a longer-term policy signal that still requires continued observation. It does not yet justify broad conclusions beyond the stated scope, but it does justify closer operational preparation by affected businesses.
Basis of This Article and What Still Needs Verification
This article is based on the user-provided news title, event date, and event summary. In coverage of developments of this type, commonly relevant source categories may include official agency announcements, company notices, industry association updates, reports by authoritative media, and documents from standards organizations.
No specific official source link was provided in the input, so the exact original publication path still needs to be continuously verified. Follow-up attention should remain on any later FDA clarification related to implementation details, the scope of the AMIFT pilot, and any further guidance affecting filing practice for covered equipment.