
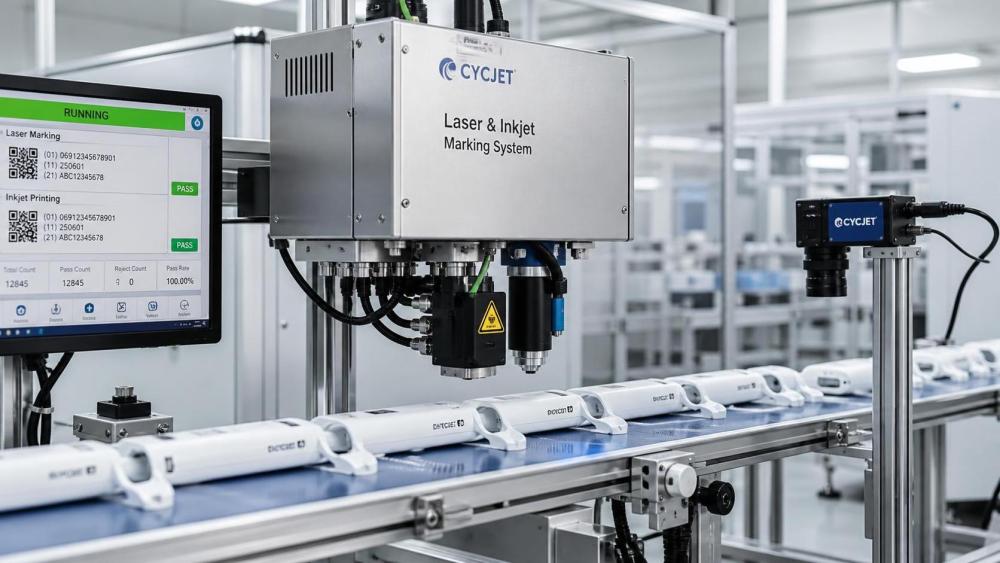
The U.S. FDA has released a draft revision to 21 CFR Part 830 that would tighten traceability requirements for medical devices entering the U.S. market. Based on the information provided, the proposed rule would require UDI-DI markings created by fiber laser markers, UV laser markers, and continuous inkjet printers on Class II and higher medical devices to support real-time verification.
In practical terms, that means the marking must be immediately readable and verifiable through systems such as Traceability Vision Systems or cloud-based parsing and validation tools. The draft also raises the bar on durability, requiring the marking to pass ISO/IEC 15415-2025 testing at grade B or above.
What the draft signals
From an industry perspective, this is not just a labeling update. It points to a more demanding compliance environment in which the mark itself is treated as part of the product’s traceability infrastructure, not only as a printed identifier. For manufacturers and exporters, that shifts attention from basic code generation to end-to-end readability, durability, and validation at the point of production.
The direct impact is likely to be felt first by Chinese equipment exporters, OEM manufacturers, and overseas distributors serving the U.S. market. If the phased rollout begins on 2027-02-01 as stated in the draft, companies will need enough lead time to check whether their marking processes, inspection systems, and downstream verification workflows can meet the new expectation consistently.
Operational pressure points
The stricter requirement may affect several parts of the supply chain at once. Marking equipment suppliers will need to focus on whether their systems can produce stable UDI-DI marks under different substrates and production conditions. Device manufacturers will need to confirm that the marks remain legible after handling, packaging, and transport. Distributors may also need to assess whether incoming goods can be validated quickly enough to avoid delays in receipt or market release.
Because the draft emphasizes real-time verification, the issue is likely to extend beyond print quality alone. It also raises questions about integration between marking hardware, inspection software, and data systems used for traceability records. Companies with fragmented workflows may face more friction than those with a unified validation process.
What to watch next
At this stage, the most relevant follow-up is the final text of the rule and how the phased implementation will be structured. For now, the safest response is to treat the draft as a clear compliance signal and begin internal review of marking durability, verification coverage, and documentation readiness.
Based on the information provided, the rule would primarily raise the compliance threshold for Class II and higher medical devices sold into the U.S. market. How widely and how quickly it changes actual procurement and export behavior will depend on the final implementation details and how strictly the verification requirement is enforced in practice.